

Here's how it works
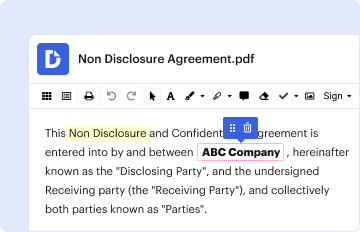
01. Edit your form online
Type text, add images, blackout confidential details, add comments, highlights and more.
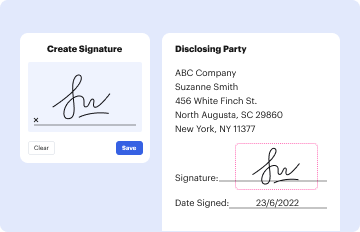
02. Sign it in a few clicks
Draw your signature, type it, upload its image, or use your mobile device as a signature pad.
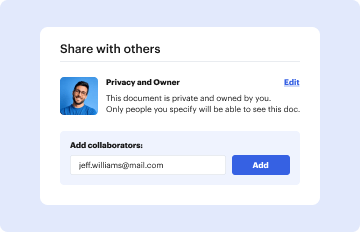
03. Share your form with others
Send it via email, link, or fax. You can also download it, export it or print it out.
How to use or fill out 510(k) Summary 82012 8 21 CFR 807 - accessdata fda with our platform
- Click ‘Get Form’ to open the document in the editor.
- Begin by filling out the submitter and manufacturing site information. Include details such as the company name, address, telephone number, and contact person.
- Next, provide device information including the trade name, classification name, product code, and regulatory class. Ensure accuracy for compliance.
- In the predicate devices section, list any devices that your product is substantially equivalent to. This is crucial for regulatory approval.
- Describe the device's intended use clearly. Specify how it operates and its purpose in medical procedures.
- Complete the comparison of technological characteristics by detailing how your device aligns with existing products in terms of functionality and safety.
- Finally, summarize non-clinical tests conducted on your device to demonstrate compliance with FDA standards.
Start using our platform today to streamline your document editing and submission process for free!
Fill out 510(k) Summary 82012 8 21 CFR 807 - accessdata fda online
It's free
It's free
Got questions?
We have answers to the most popular questions from our customers. If you can't find an answer to your question, please contact us.
Contact us
What are the requirements for 510k clearance?
The FDA 510k submission requirements include extensive technical documentation such as proposed labeling, sterilization and shelf life data, biocompatibility research, animal, clinical and bench performance testing and other technical evidence needed to establish substantial equivalence and receive FDA 510k clearance.
How long does FDA 510k clearance take?
Once the FDA have received your submission, the FDA will officially start the clock on your decision timeline. At the time of writing the FDA states that a traditional 510(k) decision should be provided within 90 days.
What is 21 CFR Part 807 FDA?
Owners or operators of establishments that are involved in the production and distribution of medical devices intended for use in the U.S. are required to register annually with the FDA. This process is known as establishment registration (Title 21 CFR Part 807).
What is FDA 510k approved?
The FDAs 510(k) submission process is, in a nutshell, a clearance process which every company wanting to bring a medium-risk medical device to the American market must pass through. Almost half of all medical devices used in the United States every day will have passed through the 510(k) route.
What is the difference between FDA and FDA 510k?
A 510(k) is a premarket submission made to FDA to demonstrate that the device to be marketed is as safe and effective, that is, substantially equivalent, to a legally marketed device (section 513(i)(1)(A) FDC Act).
Security and compliance
At DocHub, your data security is our priority. We follow HIPAA, SOC2, GDPR, and other standards, so you can work on your documents with confidence.
Learn more


People also ask
What does FDA granted 510 K clearance mean?
The 510(k) clearance process involves a comprehensive review of safety and performance data for the device, which may include scientific, non-clinical, and clinical data, as appropriate, to determine if a new device is substantially equivalent to a device that is already on the market (that is, a predicate device).